Proksee News
Proksee’s updated label system gives you more control over how feature labels appear on your maps. Choose between Classic and Adaptive placement, adjust label density, and access the main settings through a new popup beside your map.
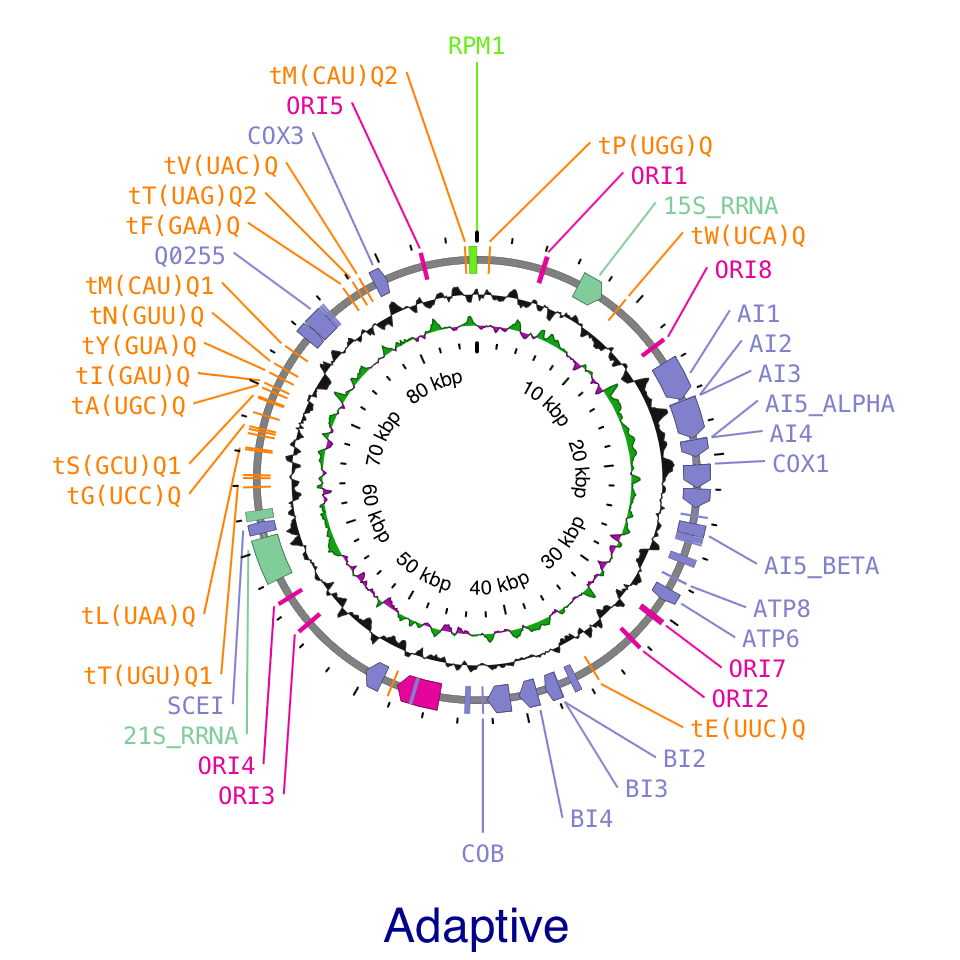
Adaptive adjusts label placement to the density of features and the available space. At low density, labels follow an oval around a full circular map. As density increases, crowded regions reorganize to fit more labels while less crowded regions can retain a simpler arrangement. Smooth transitions during panning and zooming make labels easier to follow as their positions change.
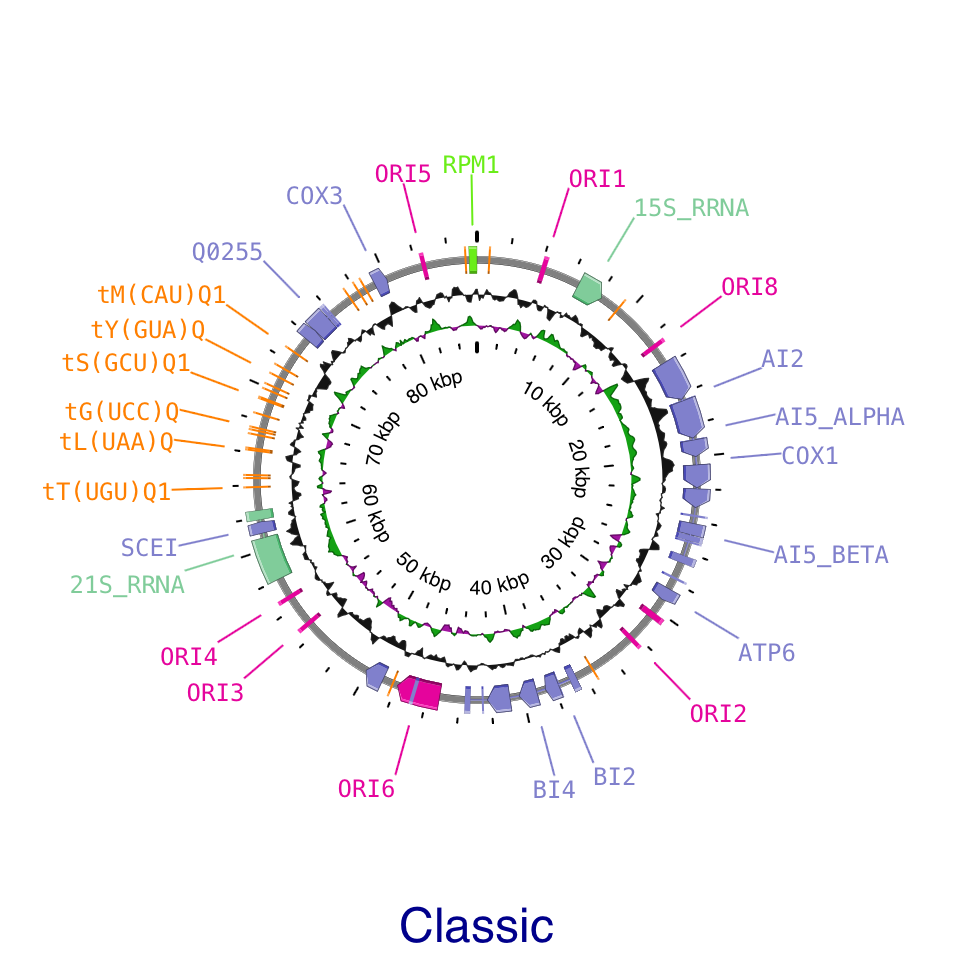
Classic uses straight leader lines and arranges labels in outward levels, providing a familiar layout for both circular and linear maps.
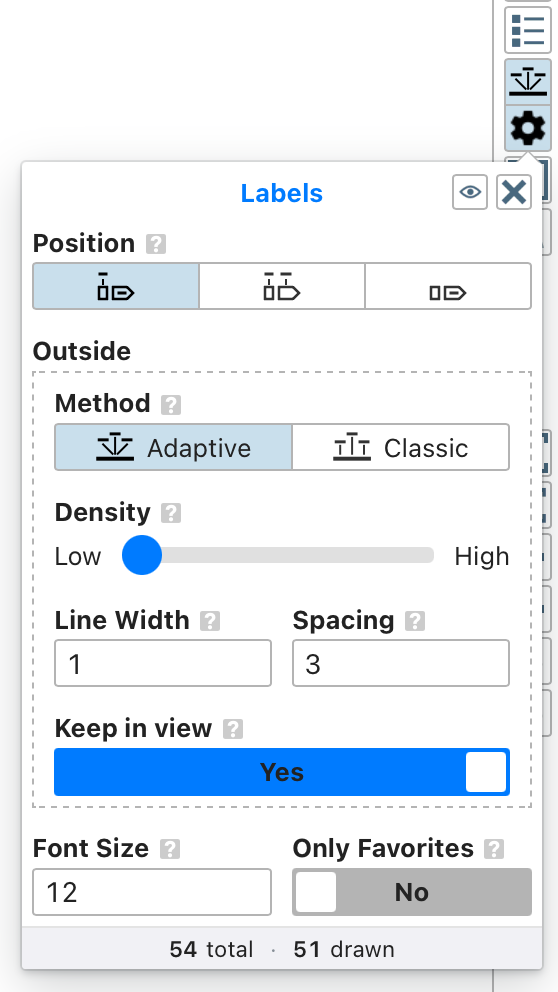
Both methods share a Density slider. Move toward Low for a cleaner layout or toward High to display more labels. You can also keep labels within the visible map area and adjust their spacing, font size, and leader line width.
Click the gear beneath the label visibility button to open the new settings popup. It provides quick access to the main label controls and shows the total number of labels and the number currently drawn. Click the Labels title to open the full settings in the sidebar.
Existing maps are updated automatically when opened: Default placement becomes Classic, and Angled placement becomes Adaptive. Adaptive is the default for new maps, and you can switch between the two methods at any time.
A new tool has been added to Proksee:
Genome Identity Matrix
- Compare two or more complete or draft microbial genome assemblies with FastANI.
- Explore directional average nucleotide identity (ANI), distance, and alignment-fraction matrices.
- Group similar genomes with average-linkage clustering and matching dendrograms.
- Inspect FastANI fragment mappings for individual query–reference comparisons.
- Download matrix tables, figures, pairwise views, and summary metadata for downstream review.
FastANI was developed by Chirag Jain and colleagues for fast, whole-genome ANI estimation. The method and its large-scale application are described in their Nature Communications paper, High throughput ANI analysis of 90K prokaryotic genomes reveals clear species boundaries.
Genome Identity Matrix packages all directional FastANI comparisons into one interactive report. Users can switch among ANI, distance, and alignment fraction; view genomes in input or clustered order; add dendrograms; search genome labels; and inspect exact values for each query–reference cell. The displayed matrix can be exported as SVG, PNG, or ordered CSV.
The examples below use the tool's five-genome diagnostic data set. The 600 kb assembly, close_a, and close_b genomes produce dense collinear mappings. The 600 kb rearranged genome retains sixteen 30 kb blocks in a changed order, mixes in four unrelated blocks, and is stored as twenty FASTA records. The distant genome provides comparisons for which FastANI reports no value.
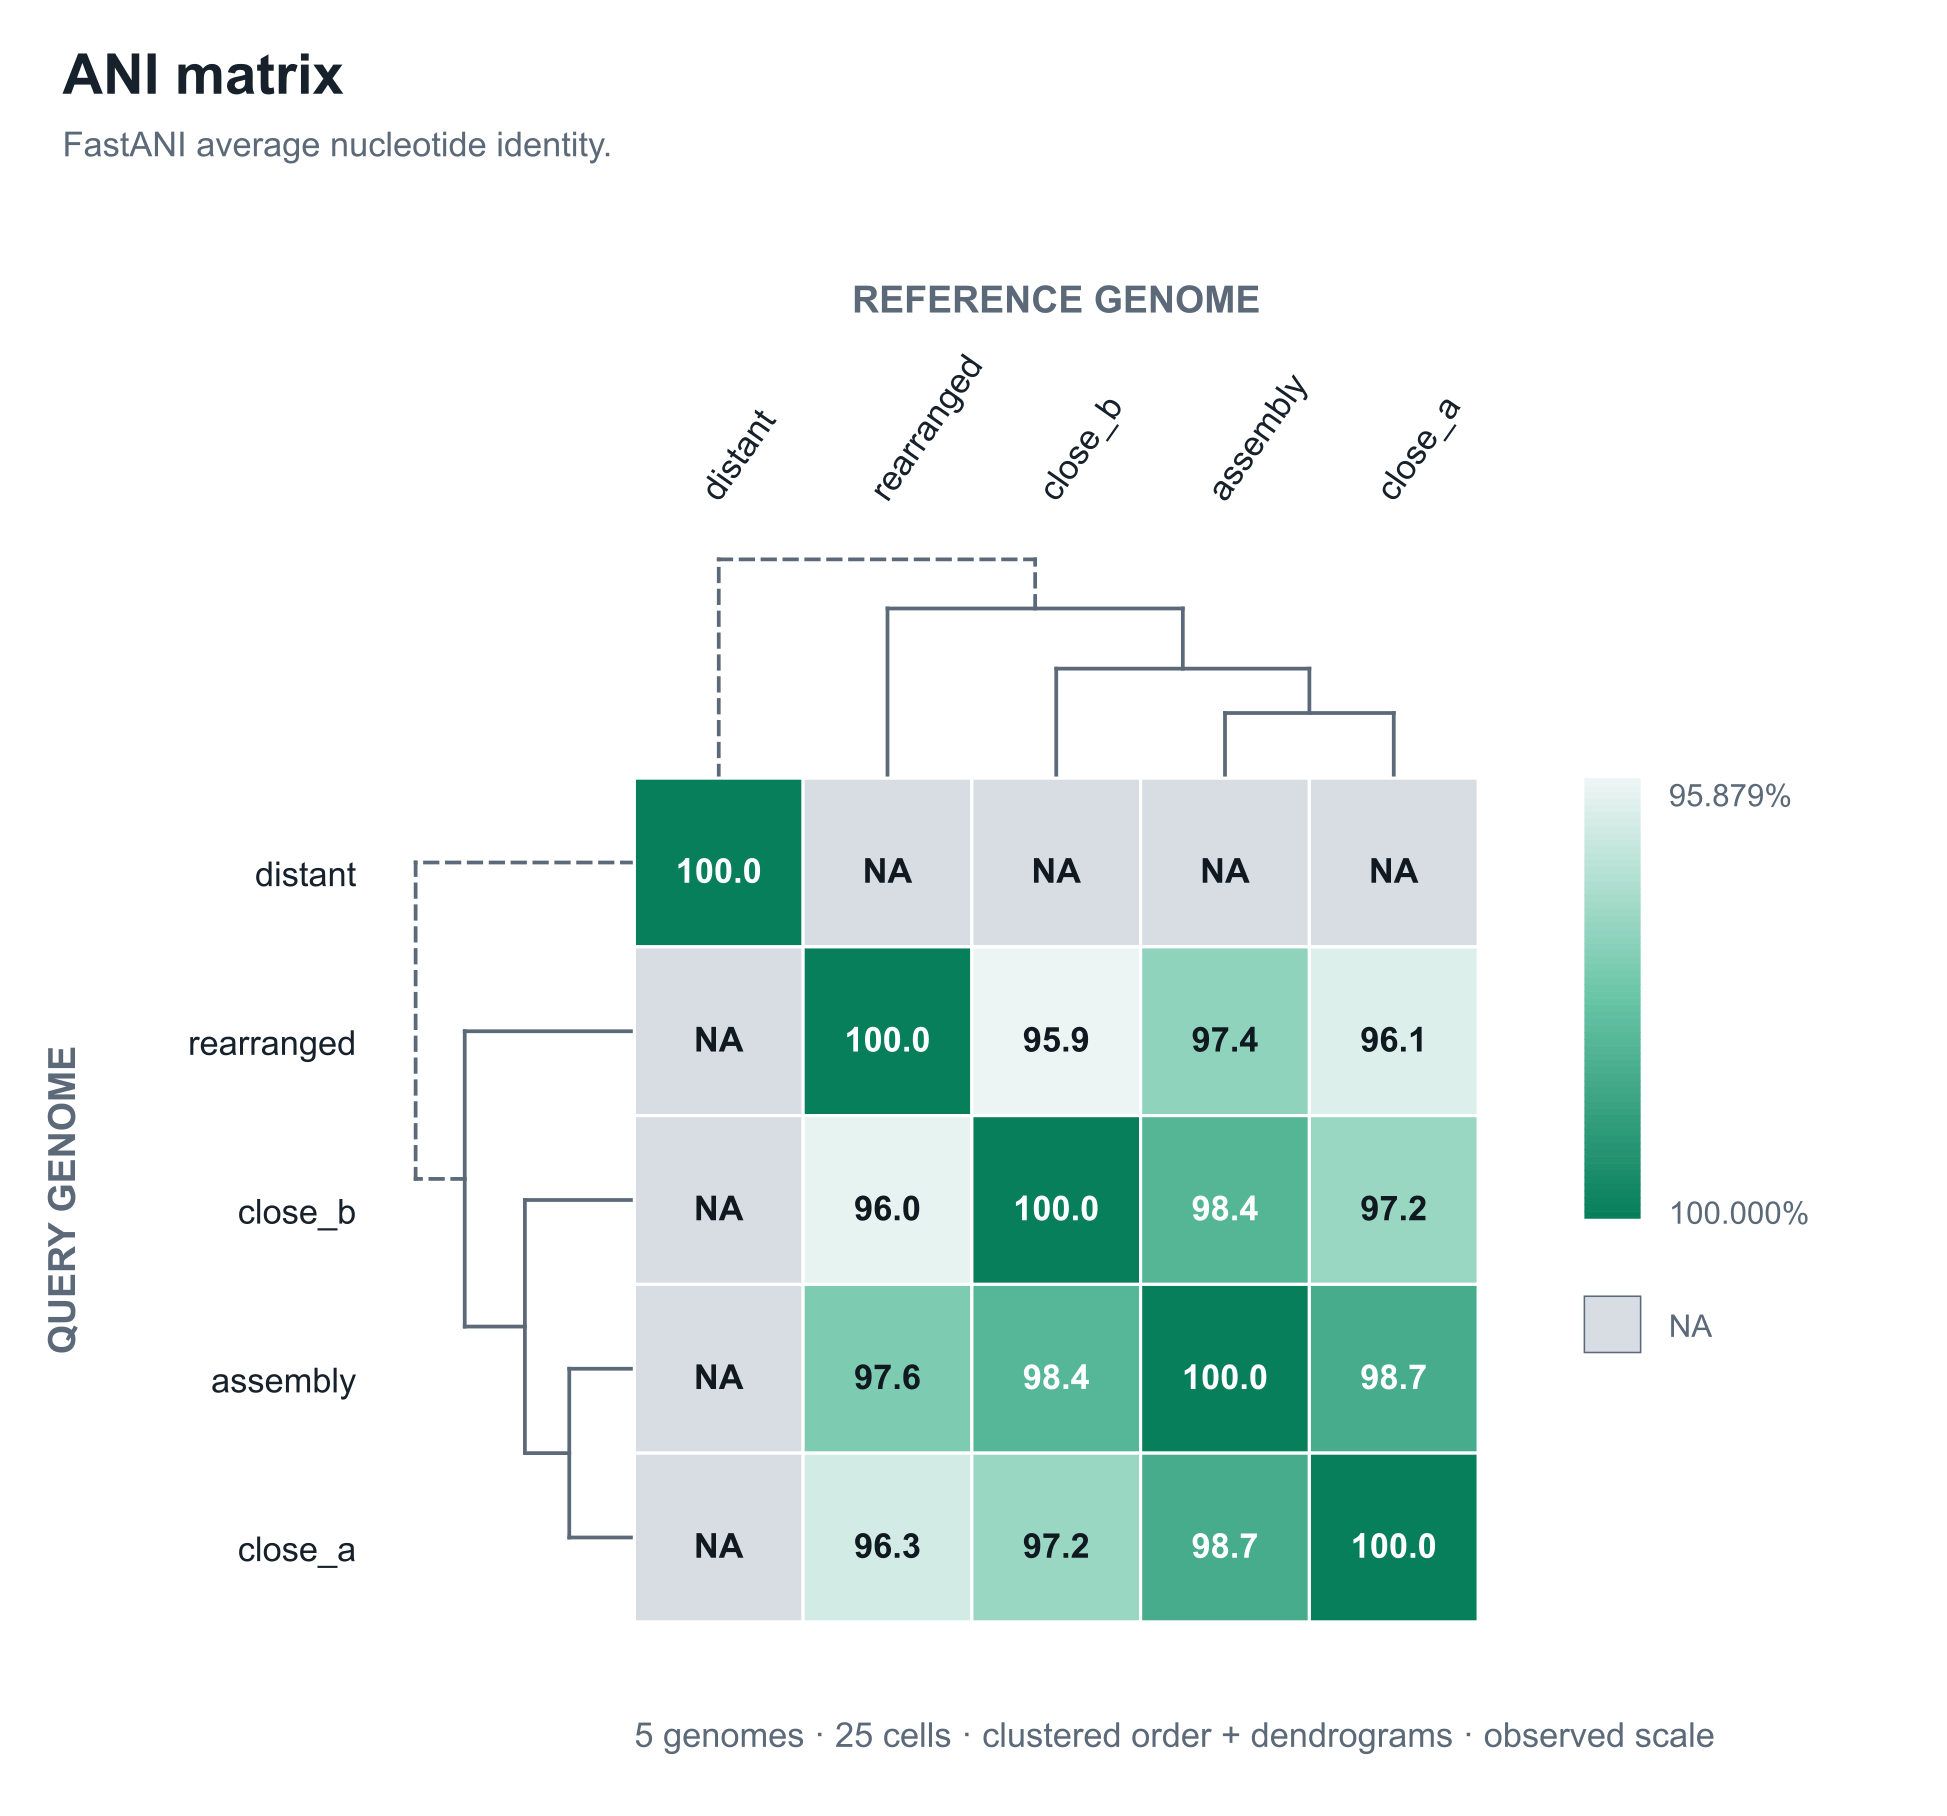
The dendrograms use average-linkage clustering on 100 − ANI, combining available results from both comparison directions. Solid joins are supported by FastANI distances; dashed joins indicate that no FastANI value was available between the connected clusters.
Selecting a matrix cell opens its directional FastANI fragment-mapping view. The reference genome is shown on top and the query genome below on one shared base-pair scale. A reference-position colour is copied to each linked query block, so smooth colour order supports conserved placement while reordered colours and crossing ribbons reveal changed sequence order or placement. Uncoloured intervals lack a plotted FastANI mapping, and darker ribbons represent higher fragment identity.
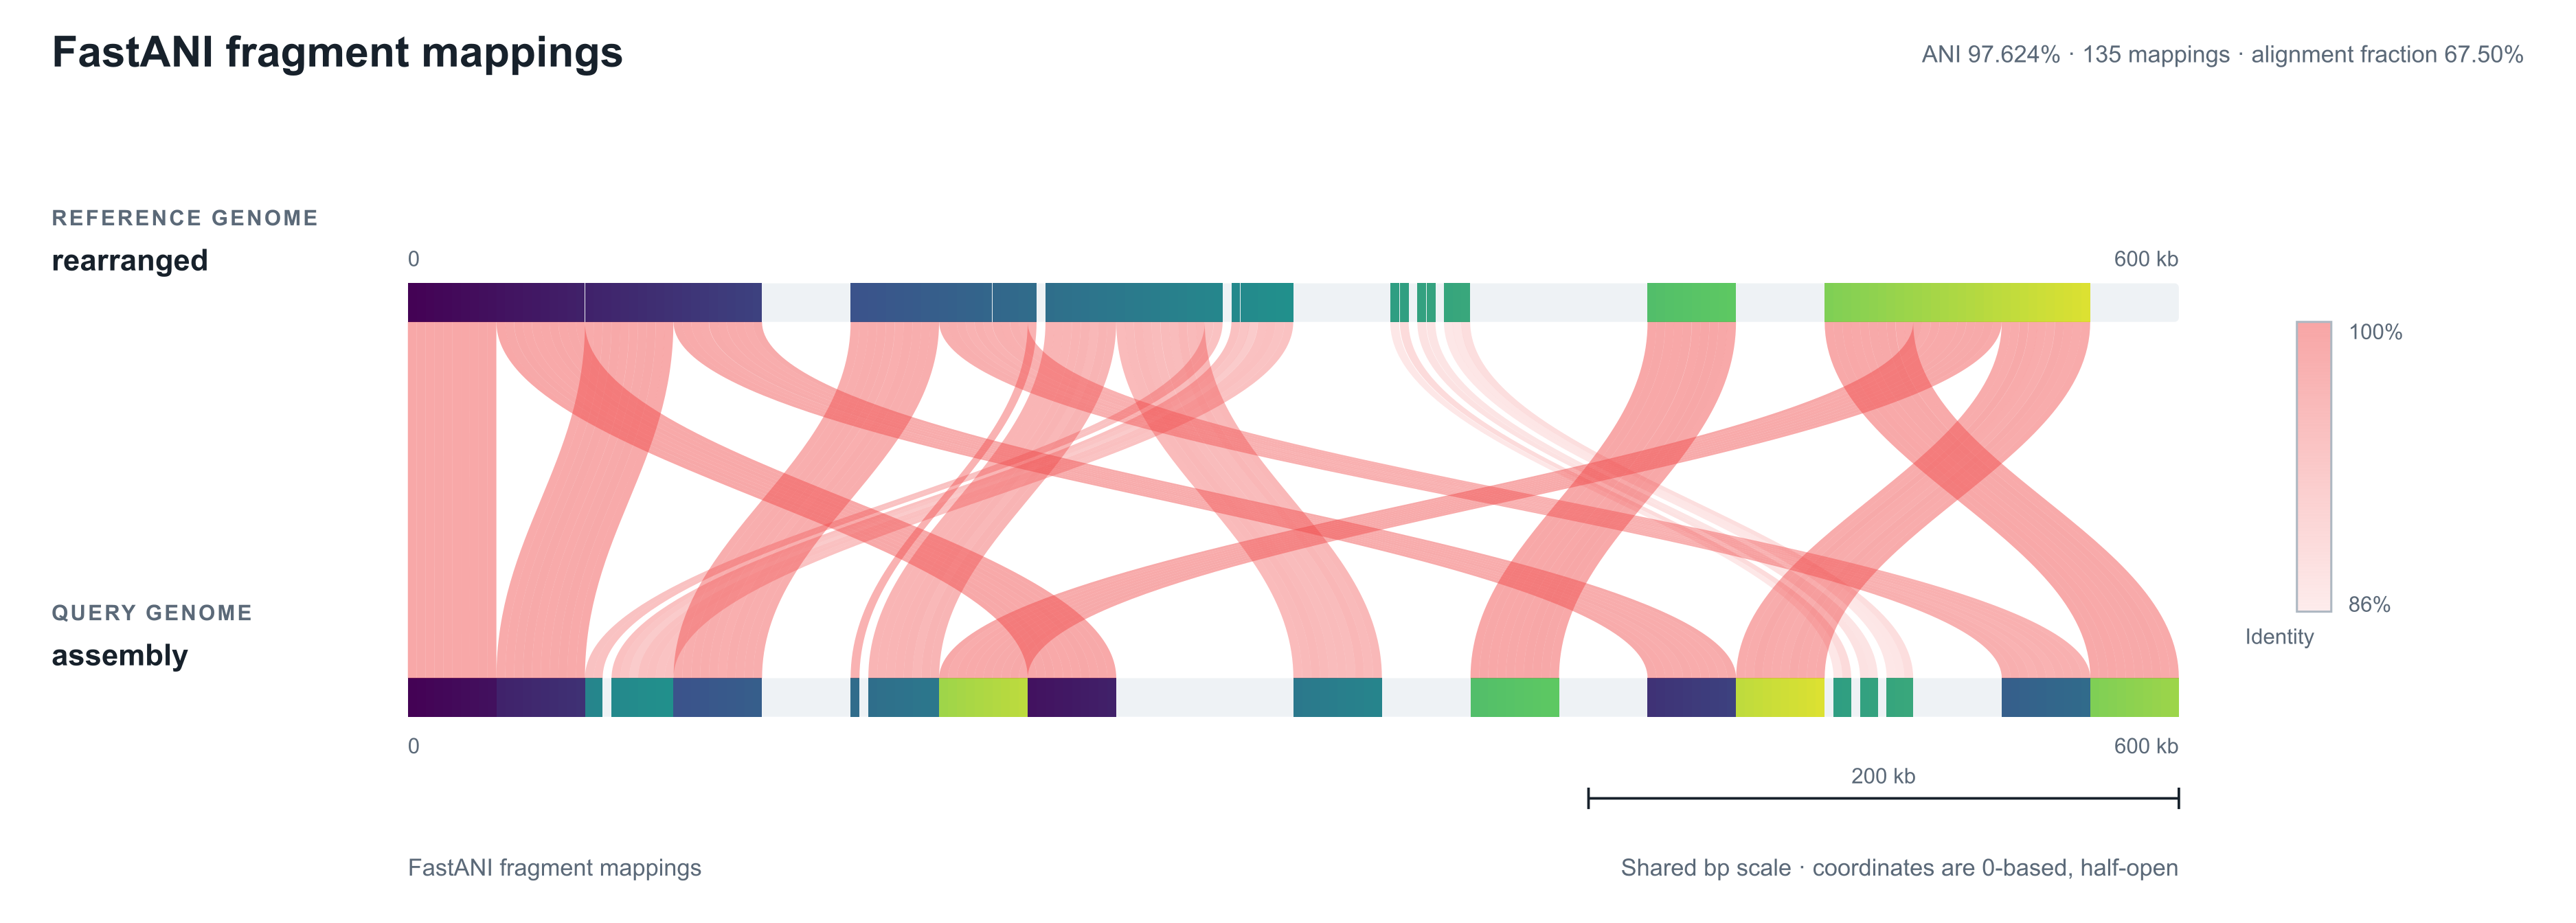
Genome Identity Matrix runs FastANI separately in both directions for every genome pair. The two results remain visible in opposite matrix cells and in separate pairwise views; the displayed ANI and alignment fraction values are not averaged. Because alignment fraction is calculated relative to the query genome, the two directions can differ. Fragmented-assembly record order can also create rearrangement-like mapping patterns, and FastANI's visual output does not report strand, so unexpected patterns are best followed with more detailed analysis.
Proksee’s latest map-viewer update brings six-frame translation, richer sequence detail, and new feature and track labels. Powered by CGView 1.9, these additions make it easier to explore your genome at different scales.
- Explore all six reading frames. Zoom into the nucleotide sequence to see amino-acid translations on both strands, with start and stop codons highlighted. Hover over a translated codon to inspect its amino acid, sequence, position, reading frame, and genetic code.
- See more sequence detail. Distinct nucleotide colors and directional boxes make individual bases easier to distinguish. Choose colored boxes, uniform boxes, or letters only, with horizontal text or text that follows a circular map.
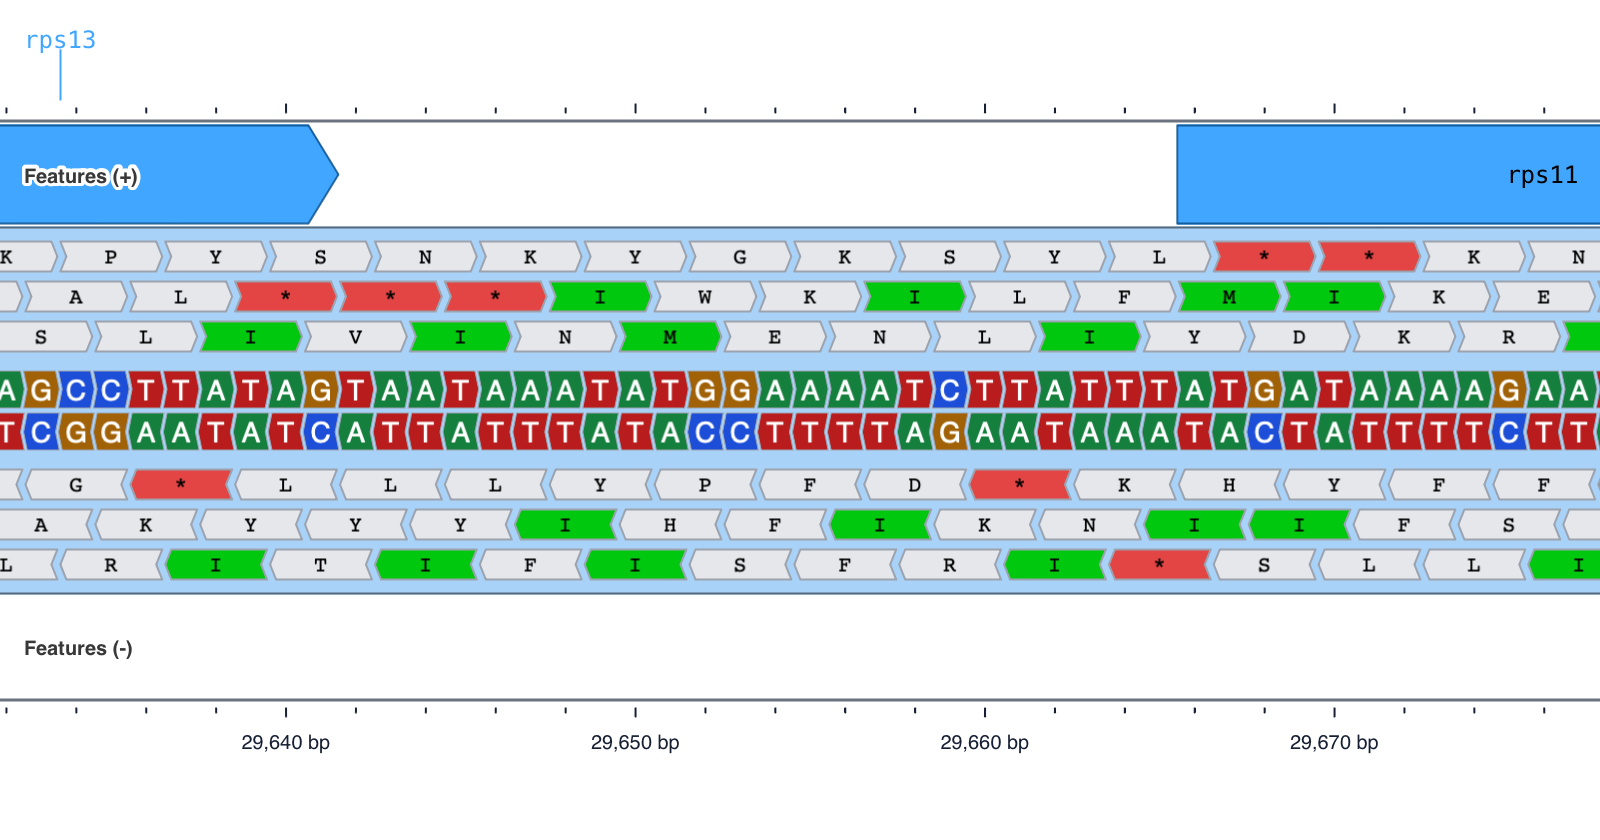
- Read feature names directly on the map. Inline labels place names inside their features. Auto placement uses inline labels where they fit and falls back to outside placement. Optional shrinking and truncation provide additional control.
- Identify tracks as you zoom in. Labels identify each track and its individual slots, helping you stay oriented while exploring a region.
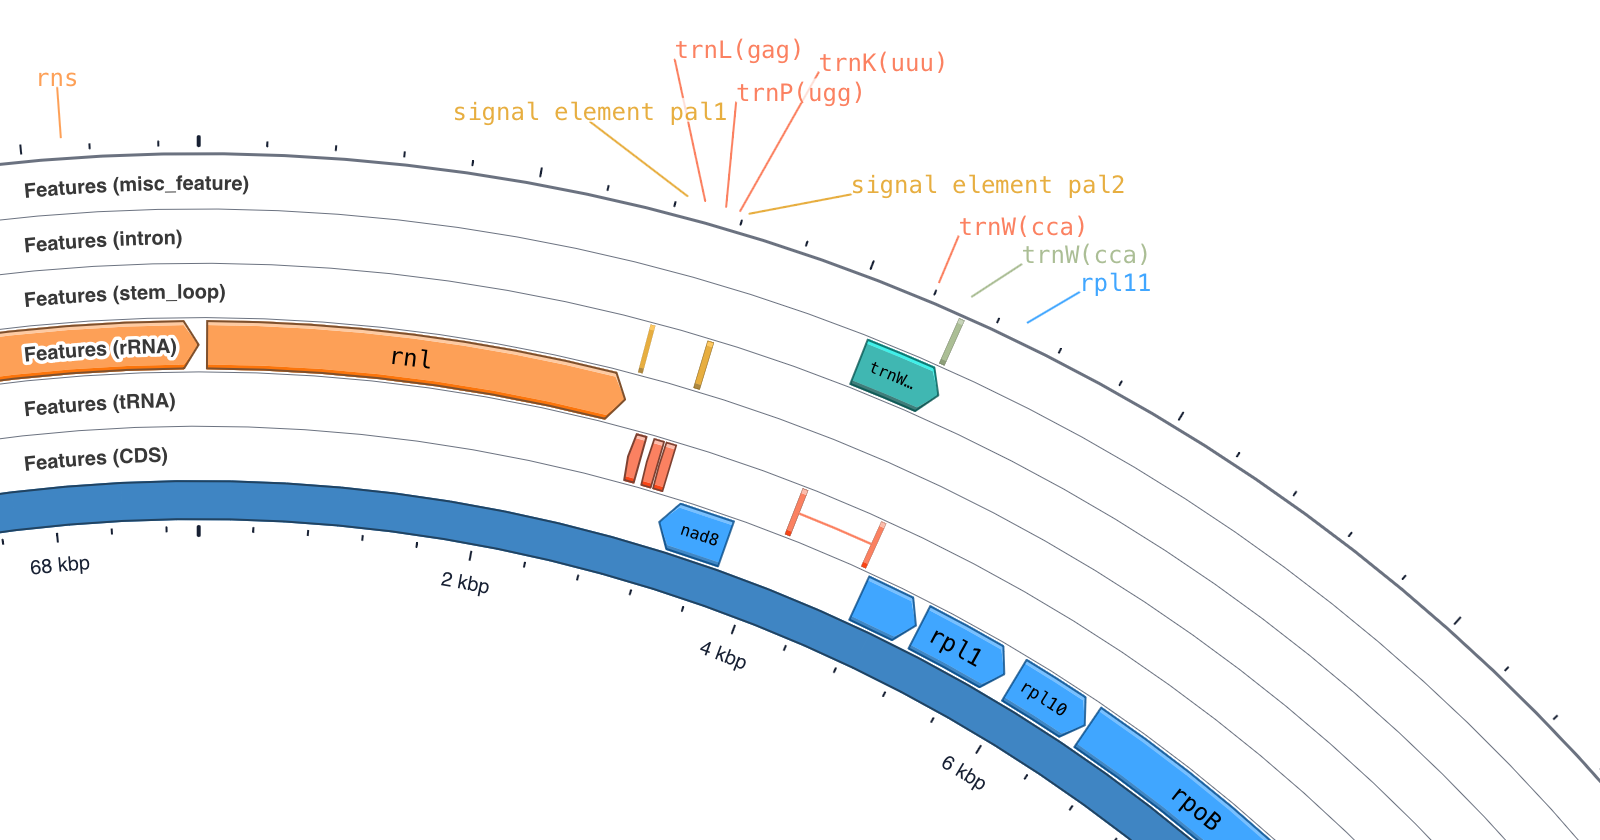
- Let feature arrows adapt to zoom. The new Auto option displays features as arcs when they are small on screen and progressively reveals arrowheads as you zoom in.
- Adjust track thickness more precisely. Fine-tune the width of track slots using relative ratios or pixel values.
- Choose where ruler labels appear. Place coordinate labels inside, outside, or on both sides of the map, or hide them. Circular maps also support labels that follow the curve.
The update also improves the contours of line plots such as GC content and GC skew, and fixes blurry arcs when viewing zoomed circular maps in Safari. Proksee now remembers your selected sidebar tabs between visits.